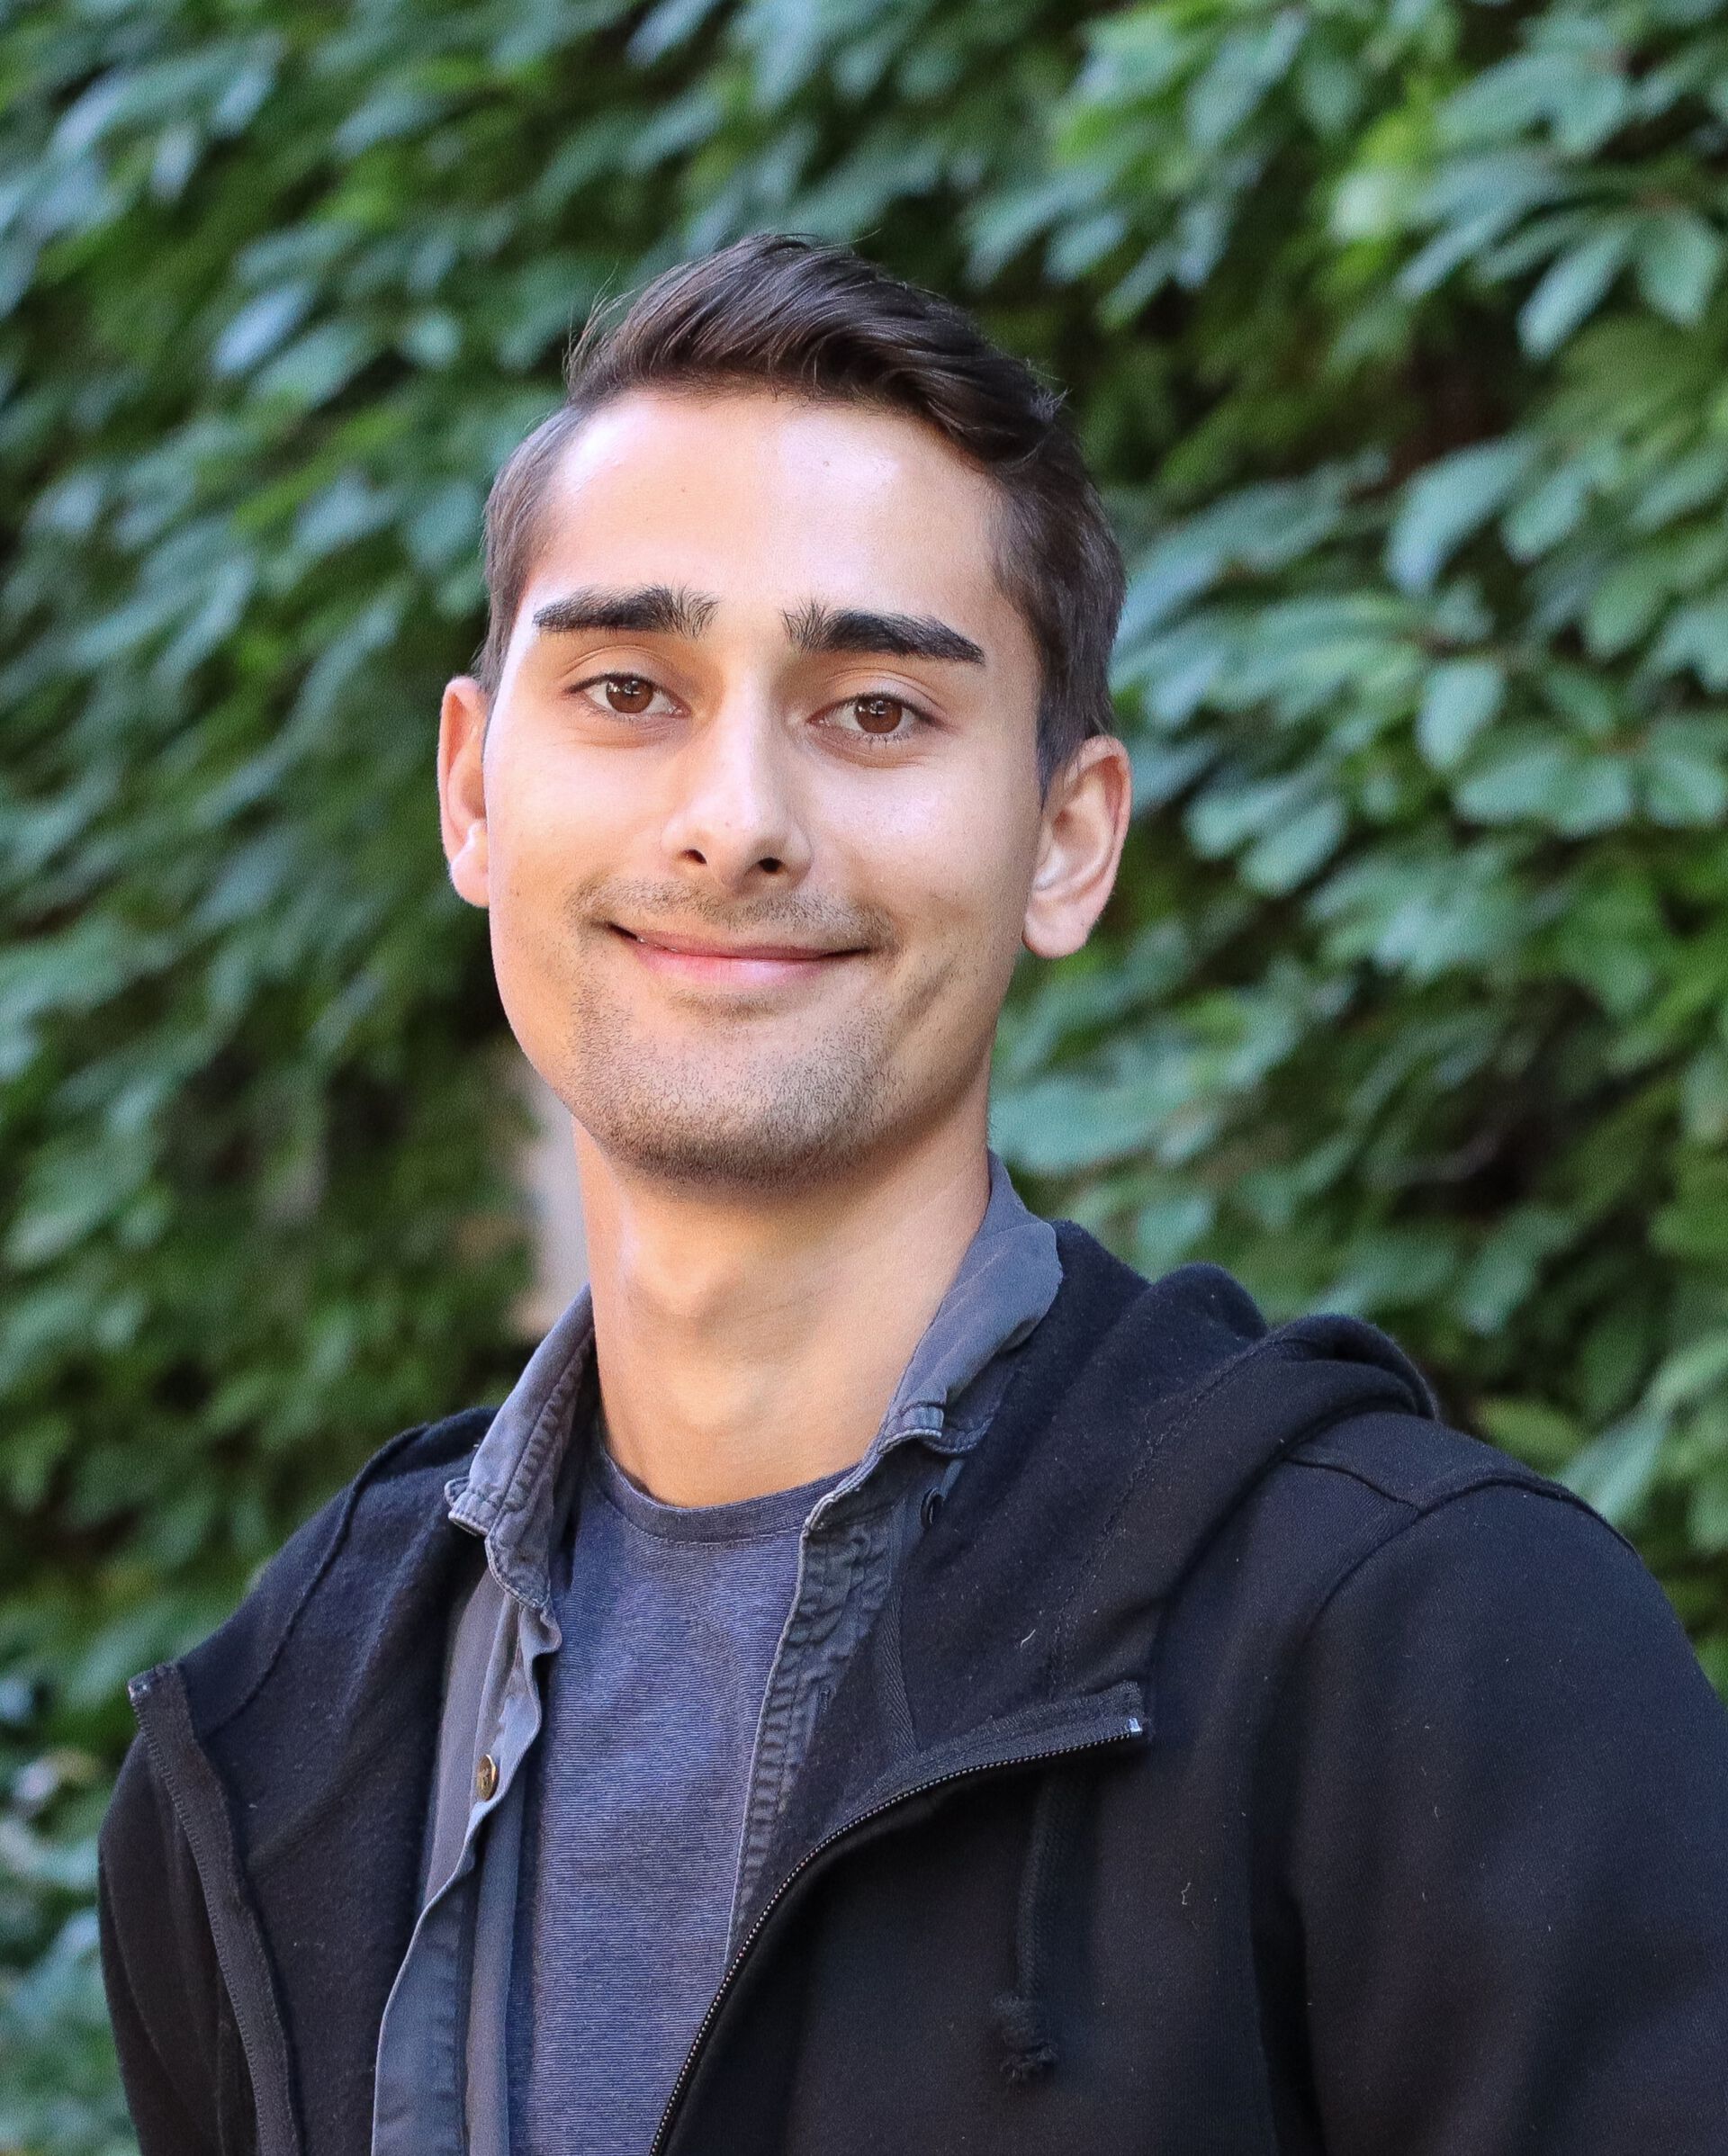
Academic interests
I am a PhD student at the CEES and a part of the project "Lacewing venom: Linking the molecular and phenotypic evolution of adaptive traits" (ERC), led by Eivind Andreas Baste Undheim. I work with the macro-evolutionary aspect of the project, studying the mechanisms by which neuropteran venom proteins arose and have diversified using a broad range of methods, including proteomics, transcriptomics, genomics, functional assays, molecular evolutionary analyses, and structural biology.
Teaching assistant
Background
I did my MSc at the CEES, working with evolutionary genomics, including population genomics and phylogenomics, of Arctic fishes.
Tags:
Evolution,
Genomics,
Venomics
Publications
-
Maurstad, Marius Filomeno; Kusy, Dominik; Undheim, Eivind Andreas Baste; Jakobsen, Kjetill Sigurd; Böhne, Astrid & Monteiro, Rita
[Show all 20 contributors for this article]
(2026).
ERGA-BGE chromosome-level genome assembly of the giant stream lacewing Osmylus fulvicephalus (Scopoli, 1763).
Open Research Europe.
6.
doi:
10.12688/openreseurope.23457.3.
Full text in Research Archive
Show summary
The giant stream lacewing, Osmylus fulvicephalus (Scopoli, 1763), is a widespread European species belonging to the insect order Neuroptera. Its cryptic larvae are predators found at the banks of streams and smaller rivers where they use their piercing, lance-shaped stylets to inject venom into their arthropod prey. Here, we present the reference genome of the giant stream lacewing as a crucial resource for uncovering the genetic basis of venom evolution in Neuroptera. The chromosome-level genome encompasses 674.7 Mb and is composed of 60 contigs and 24 scaffolds where 99.2% of the assembly is distributed among the 6 contiguous chromosomal pseudomolecules and two sex chromosomes (X and Y). Contig and scaffold N50 have a value of 51.5 Mb and 116.2 Mb, respectively. This reference genome is the first genomic resource from the family of lance lacewings, providing valuable data for clarifying the phylogenetic placement of the family Osmylidae within Neuroptera.
-
Araya, Robin Aasegg; Maurstad, Marius Filomeno; Wilson, David T; Rash, Lachlan D; Mobli, Mehdi & Jakobsen, Kjetill Sigurd
[Show all 7 contributors for this article]
(2026).
Ancient origin and dynamic evolution of bivalent spider toxins.
Molecular Biology and Evolution (MBE).
ISSN 0737-4038.
43(5).
doi:
10.1093/molbev/msag076.
Full text in Research Archive
Show summary
Bivalent peptide toxins comprising 2 cysteine-rich domains have evolved from single-domain precursors on multiple occasions in animal venoms, resulting in enhanced molecular target selectivity and avidity. Although bivalent toxins are emerging as prevalent in animal venoms, the genomic and evolutionary processes driving the transitions between single- and multi-domain architectures remain poorly understood. Here, we investigated the evolution of bivalent inhibitor cystine knot (ICK) toxins in spider venom. We first generated a genome assembly of the tree-dwelling funnel-web spider Hadronyche cerberea, revealing a massive expansion of ICK toxin-encoding genes, including the bivalent π-hexatoxin-Hc1a. All ICK toxin genes share a conserved 3-exon structure, flanked by transposable elements (TEs) that may have facilitated gene expansion. This gene structure is shared by the Hc1a subfamily, where the entire mature bivalent toxin is encoded by the third exon. Leveraging de novo transcriptome assemblies from 86 spider species along with venom proteomic data, we show that bivalency in the Hc1a subfamily is of ancient origin and evolved via intra-exonic duplication not involving introns. This was followed by domain expansion and recurrent domain losses mediated by point mutations, deletions, and unequal crossing-over facilitated by high interdomain sequence similarity. In contrast, the bivalent toxin DkTx from Cyriopagopus schmidti is confined to a small group of tarantulas, where it appears to have evolved once, with subsequent domain losses potentially linked to TE activity. Our findings reveal that singular events of domain duplication can give rise to complex, asymmetrical evolutionary trajectories shaped by gene instability and selective retention of functional domains.
-
Hoff, Siv Nam Khang; Maurstad, Marius Filomeno; Tørresen, Ole K.; Araya, Robin Aasegg; Berg, Paul R. & Præbel, Kim
[Show all 8 contributors for this article]
(2026).
Rapid genome modifications including chromosomal fusions and large-scale inversions are key features in Arctic codfish species.
Genome Biology.
ISSN 1474-7596.
27.
doi:
10.1186/s13059-026-03975-6.
Full text in Research Archive
Show summary
Genome evolvability involves activation of transposable elements (TEs) that result in novel genomic rearrangements, including translocations, deletions, duplications, as well as larger structural reorganizations, such as chromosomal inversions and fusions. These genomic modifications contribute to raw genetic variability in which selection can act upon, and thus, promote local adaptation. Results Using a comparative genomics framework combined with the generation of six chromosome-level gadid reference genomes, including the cold-water adapted polar cod ( Boreogadus saida ) and Arctic cod ( Arctogadus glacialis ), we uncover an array of larger and smaller chromosomal reorganizations within this lineage. For the two Arctic codfishes, we detect lineage-specific chromosomal fusions, i.e., five in polar cod vs. eight in Arctic cod, resulting in a reduced number of chromosomes found to be associated with their geographical distribution. For the same species, we identify a high number of partly overlapping chromosomal inversions where a majority (especially within breakpoint regions) display an overrepresentation of specific TE families, accompanied by signatures of high degree of conservation. We further demonstrate involvement of miniature inverted-repeat transposable elements (MITEs) in the expansion of the well-known antifreeze glycoprotein genes in polar cod, and that the three gene clusters were localized in association with chromosomal rearrangements on chromosomes 1, 2, and 14. Conclusions We characterize how the Gadidae lineage has undergone massive genomic modifications — potentially via activation of TEs — throughout their evolutionary history, and particularly for the more Arctic species. These genomic reorganizations have likely played an important role in divergence processes and adaptation to freezing environmental conditions.
-
Maurstad, Marius Filomeno; Hoff, Siv Nam Khang; Cerca, José; Ravinet, Mark; Bradbury, Ian & Jakobsen, Kjetill Sigurd
[Show all 8 contributors for this article]
(2025).
Reference genome bias in light of species-specific chromosomal reorganization and translocations.
Genome Biology.
ISSN 1465-6906.
26(1),
p. 1–26.
doi:
10.1186/s13059-025-03761-w.
Full text in Research Archive
Show summary
Abstract Background Whole-genome sequencing efforts, have during the past decade, unveiled the central role of genomic rearrangements—such as chromosomal inversions—in evolutionary processes, including local adaptation in a wide range of taxa. However, employment of reference genomes from distantly or even closely related species for mapping and the subsequent variant calling can lead to errors and/or biases in the datasets generated for downstream analyses. Results Here, we capitalize on the recently generated chromosome-anchored genome assemblies for Arctic cod ( Arctogadus glacialis ), polar cod ( Boreogadus saida ), and Atlantic cod ( Gadus morhua ) to evaluate the extent and consequences of reference bias on population sequencing datasets (approx. 15–20 × coverage) for both Arctic cod and polar cod. Our findings demonstrate that the choice of reference genome impacts the mapping statistics, including mapping depth and mapping quality, as well as core population genetic estimates, such as heterozygosity levels, nucleotide diversity (π), and cross-species genetic divergence (D XY ). Furthermore, using a more distantly related reference genome can lead to inaccurate detection and characterization of chromosomal inversions, i.e., in terms of size (length) and location (position), due to inter-chromosomal reorganizations between species. Additionally, we observe that some of the verified species-specific inversions are split across multiple genomic regions when mapped against a heterospecific reference. Conclusions Inaccurate identification of chromosomal rearrangements as well as biased population genetic measures could potentially lead to erroneous interpretation of species-specific genomic diversity, impede the resolution of local adaptation, and thus, impact predictions of their genomic potential to respond to climatic and other environmental perturbations.
-
Maurstad, Marius Filomeno; Ramiro, Iris Bea; Øyen, Jan Philip; Sombke, Andy; Büsse, Sebastian & Nachtigall, Pedro Gabriel
[Show all 8 contributors for this article]
(2025).
The green lacewing venom system and the complex mechanisms underlying its evolution.
Molecular Biology and Evolution (MBE).
ISSN 0737-4038.
43.
doi:
10.1093/molbev/msaf326.
Full text in Research Archive
Show summary
Abstract Venom has independently evolved across many lineages, yet relatively few have been studied in detail, particularly among insects. Of these, Neuroptera (lacewings, antlions and relatives) remain largely unexplored, despite being widespread with agriculturally important groups such as green lacewings. While adults are non-venomous, neuropteran larvae are ferocious predators that use pincer-like mouthparts to inject paralysing and liquefying venom to subdue and consume their prey. Here, we provide a comprehensive investigation of the venom system in Neuroptera by integrating a high-quality genome, long-read transcriptomes spanning all life stages, microCT-reconstruction of venom glands, tissue-specific expression analyses, venom proteomics, and functional assays of the common green lacewing Chrysoperla carnea. We provide a re-description of the neuropteran venom system, demonstrate the venom’s insecticidal and cytotoxic activity, and show the venom comprises diverse toxin gene families and is richer and more similar to the venom of antlions than previously proposed. We show that this toxin arsenal is the result of a multitude of evolutionary events that include co-option, recruitment following gene duplication, diversification of toxin-paralogs by gene duplication, and functional innovation of new paralogs through both small structural and large architectural changes. In addition, we find that alternative splicing of toxin genes is an important contributor to the biochemical arsenal, which is a mechanism rarely documented among venomous animals. Our results demonstrate how multiple genomic and evolutionary mechanisms together contribute to the emergence and evolution of a complex molecular trait, and provide new insights into the evolution of venom in insects.
-
Helmerson, Cecilia; Durant, Joël; Hoff, Siv Nam Khang; Brieuc, Marine Servane Ono; Berg, Paul Ragnar & Jorde, Per Erik
[Show all 14 contributors for this article]
(2025).
A century of anthropogenic perturbations impact genomic signatures of the iconic migratory Atlantic cod.
Science Advances.
11(31).
doi:
10.1126/sciadv.adp3342.
Full text in Research Archive
Show summary
Anthropogenic stressors have led to marked ecosystem perturbations, including population declines and shifts in habitat range for key marine fish species. Understanding how these changes affect genome-wide characteristics, causing long-term evolutionary responses, is still in its infancy. Genome-wide retrospective assessment of the iconic migratory Atlantic cod ( Gadus morhua ) from the Barents Sea, unraveled varying degree of admixture with the nonmigratory coastal ecotype throughout the 20th century, and intriguingly more intensified during recent decades. These genomic changes were supported by an increased number of individuals displaying the heterozygous state of the chromosomal inversion coupled to migratory behavior in Atlantic cod. Ecological models and genome-wide scans identified that some of the observed frequency shifts, coupled to neural development, metabolic, and growth regulation, covaried with intensified fisheries, reduced generation time, and ocean warming. Our results demonstrate how anthropogenic perturbations impact the dynamics between two well-known ecotypes of Atlantic cod and thus, their genomic signatures, with potential implications for future management programs.
-
Seldeslachts, Andrea; Maurstad, Marius Filomeno; Øyen, Jan Philip; Undheim, Eivind Andreas Baste; Peigneur, Steve & Tytgat, Jan
(2024).
Exploring oak processionary caterpillar induced lepidopterism (Part 1): unveiling molecular insights through transcriptomics and proteomics.
Cellular and Molecular Life Sciences (CMLS).
ISSN 1420-682X.
81.
doi:
10.1007/s00018-024-05330-z.
Full text in Research Archive
Show summary
Lepidopterism, a skin inflammation condition caused by direct or airborne exposure to irritating hairs (setae) from processionary caterpillars, is becoming a significant public health concern. Recent outbreaks of the oak processionary caterpillar (Thaumetopoea processionea) have caused noteworthy health and economic consequences, with a rising frequency expected in the future, exacerbated by global warming promoting the survival of the caterpillar. Current medical treatments focus on symptom relief due to the lack of an effective therapy. While the source is known, understanding the precise causes of symptoms remain incomplete understood. In this study, we employed an advanced method to extract venom from the setae and identify the venom components through high-quality de novo transcriptomics, venom proteomics, and bioinformatic analysis. A total of 171 venom components were identified, including allergens, odorant binding proteins, small peptides, enzymes, enzyme inhibitors, and chitin biosynthesis products, potentially responsible for inflammatory and allergic reactions. This work presents the first comprehensive proteotranscriptomic database of T. processionea, contributing to understanding the complexity of lepidopterism. Furthermore, these findings hold promise for advancing therapeutic approaches to mitigate the global health impact of T. processionea and related caterpillars.
-
Helmerson, Cecilia; Weist, Peggy; Brieuc, Marine Servane Ono; Maurstad, Marius Filomeno; Schade, Franziska Maria & Dierking, Jan
[Show all 14 contributors for this article]
(2023).
Evidence of hybridization between genetically distinct Baltic cod stocks during peak population abundance(s).
Evolutionary Applications.
16(7),
p. 1359–1376.
doi:
10.1111/eva.13575.
Full text in Research Archive
Show summary
Range expansions can lead to increased contact of divergent populations, thus increasing the potential of hybridization events. Whether viable hybrids are produced will most likely depend on the level of genomic divergence and associated genomic incompatibilities between the different entities as well as environmental conditions. By taking advantage of historical Baltic cod (Gadus morhua) otolith samples combined with genotyping and whole genome sequencing, we here investigate the genetic impact of the increased spawning stock biomass of the eastern Baltic cod stock in the mid 1980s. The eastern Baltic cod is genetically highly differentiated from the adjacent western Baltic cod and locally adapted to the brackish environmental conditions in the deeper Eastern basins of the Baltic Sea unsuitable for its marine counterparts. Our genotyping results show an increased proportion of eastern Baltic cod in western Baltic areas (Mecklenburg Bay and Arkona Basin)—indicative of a range expansion westwards—during the peak population abundance in the 1980s. Additionally, we detect high frequencies of potential hybrids (including F1, F2 and backcrosses), verified by whole genome sequencing data for a subset of individuals. Analysis of mitochondrial genomes further indicates directional gene flow from eastern Baltic cod males to western Baltic cod females. Our findings unravel that increased overlap in distribution can promote hybridization between highly divergent populations and that the hybrids can be viable and survive under specific and favourable environmental conditions. However, the observed hybridization had seemingly no long-lasting impact on the continuous separation and genetic differentiation between the unique Baltic cod stocks.
-
Pinninti, Likith Reddy; Maurstad, Marius Filomeno; Hoff, Siv Nam Khang; Kristensen, Torstein; Noble, Leslie Robert & Jentoft, Sissel
[Show all 7 contributors for this article]
(2023).
The complete mitochondrial genome of the Atlantic spiny lumpsucker Eumicrotremus spinosus (Fabricius, 1776).
Mitochondrial DNA Part B: Resources.
8(3),
p. 364–367.
doi:
10.1080/23802359.2023.2184649.
Full text in Research Archive
Show summary
The complete mitogenome of the Atlantic spiny lumpsucker (Eumicrotremus spinosus) was generated using the PacBio Sequel II HiFi sequencing platform. The mitogenome assembly has a length of 19,281 bp and contains 13 protein-coding sequences, 22 tRNA genes, 2 rRNA genes, one control region containing the D-loop (2383 bp) and a duplicate control region (1133 bp) Phylogenetic analysis using maximum likelihood revealed that E. spinosus is closely related to the Siberian lumpsucker (E. asperrimus). The mitogenome of the spiny lumpsucker will be useful in population genomics and systematic studies of Cyclopteridae, Liparidae, and Cottidae.
-
Cerca, José; Maurstad, Marius Filomeno; Rochette, Nicolas C.; Rivera-Colón, Angel G.; Rayamajhi, Niraj & Catchen, Julian M.
[Show all 7 contributors for this article]
(2021).
Removing the bad apples: A simple bioinformatic method to improve loci-recovery in de novo RADseq data for non-model organisms.
Methods in Ecology and Evolution.
12(5),
p. 805–817.
doi:
10.1111/2041-210X.13562.
Full text in Research Archive
Show summary
1. The restriction site-associated DNA (RADseq) family of protocols involves digesting DNA and sequencing the region flanking the cut site, thus providing a cost and time-efficient way for obtaining thousands of genomic markers. However, when working with non-model taxa with few genomic resources, optimization of RADseq wet-lab and bioinformatic tools may be challenging, often resulting in allele dropout—that is when a given RADseq locus is not sequenced in one or more individuals resulting in missing data. Additionally, as datasets include divergent taxa, rates of dropout will increase since restriction sites may be lost due to mutation. Mitigating the impacts of allele dropout is crucial, as missing data may lead to incorrect inferences in population genetics and phylogenetics. 2. Here, we demonstrate a simple pipeline for the optimization of RADseq datasets which involves partitioning datasets into subgroups, namely by reducing and analysing the dataset at a population or species level. By running the software Stacks at a subgroup level, we were able to reliably identify and remove individuals with high levels of missing data (bad apples) likely stemming from artefacts in library preparation, DNA quality or sequencing artefacts. 3. Removal of the bad apples generally led to an increase in loci and decrease in missing data in the final datasets. 4. The biological interpretability of the data, as measured by the number of retrieved loci and missing data, was considerably increased.
View all works in NVA
-
-
-
-
Renner, Angelika H.H.; Gradinger, Rolf; Altenburger, Andreas; Amdahl, Pernille; Barstein, Karoline & Duarte, Pedro
[Show all 31 contributors for this article]
(2022).
JC3 Winter gaps cruise: Cruise Report.
Septentrio Academic Publishing.
Full text in Research Archive
Show summary
The Nansen Legacy (NL) JC3 cruise (19.02.-11.03.2022) aimed to fill regional, temporal and scientific gaps following the earlier NL cruises, in particular during the winter-to-spring transition and in the northern part of the NL transect to the Nansen Basin. Joint physical, chemical, and biological sampling and experiments for new technology addressed aims of RF1, RF2, RF3 and RA-C.
-
Jones, Elizabeth Marie; Reigstad, Marit; Espinel, Nadjejda; Gjerland, Audun; Goraguer, Lucie Hélène Marie & Hodal, helene
[Show all 15 contributors for this article]
(2022).
JC2-1 Joint cruise part 1 2021 : Cruise Report.
Septentrio Academic Publishing.
Full text in Research Archive
Show summary
The Nansen Legacy Joint Cruise 2, part 1 (JC2-1) 12-29 July 2021, continued the investigation of the interannual variability during the late summer season. At the same time will the cruise provide a late summer reference for the seasonal investigation that was separated to late summer and polar night 2019, and winter and spring 2021. The transect represents an environmental gradient going through the northern Barents Sea, and included 7 process stations (P1-P7) lasting 10-39 hrs. Additional CTD stations (NLEG) were taken between the process stations to increase the hydrographic resolution on the transect. The work started at 76°N at the open Atlantic Water dominated station P1, was sea ice covered from station P4 at 79 45.00 °N and included deep water stations at the P7 station at 82°N in the Nansen Basin.
The program included measurements and sampling from the atmosphere, sea ice, ocean and sea floor. Data collected includes several disciplines to map the physical environment, the chemical characterization and biological communities. Key parameters from the Nansen Legacy joint cruises were selected and included in this reduced sampling program, compared to the other joint cruises. An important aim was to characterize the northern Barents Sea to map the interannual variability, but also to see how the proceeding winter and spring conditions has impacted the environment and conditions this year. A third task is to carry out a shelf investigation that can complement the investigations in the deep Arctic Basin starting in late August. Tests and improvements of a modified pelagic trawl (Harstad) were carried out as preparations for the Arctic Basin cruise in late August 2021.
-
Maurstad, Marius Filomeno
(2022).
Identification of multiple chromosomal inversions in Arctic cod (Arctogadus glacialis) – and its putative implications for defining interspecies relationships.
Universitetet i Oslo.
Full text in Research Archive
View all works in NVA
Published
Jan. 18, 2023 3:57 PM
- Last modified
June 21, 2023 2:42 PM